
ERNICA Research Collaboration Webinar in Clinical Trials
With Dr. Fleur von Hall & Dr. Ahmed Madadi Sanjani · EUPSA/ERNICA
Part of
Esophageal Atresia 52 items
Cued at 15:00 · stops at 15:45 · press play
Educational content from recorded physician discussions — not medical advice. Talk to your (or your child's) care team about your situation.
Video
STEPS-EA trial: Esophageal Atresia
3 min · Published Nov 2023
Podcast
Journal of Pediatric Surgery Article Review: 2nd Quarter (Apr-Jun) 2025
16 min · Published Aug 2025
Video
Thoracoscopic Repair of Tracheo-esophageal Fistula Tricks: Pediatric Surgery...
19 min · Published Jul 2017
Podcast
Journal of Pediatric Surgery Article Review: 1st Quarter (Jan-Mar) 2025
16 min · Published May 2025
Video
Rigid Bronchoscopy for Diagnosis and Treatment of Proximal Pouch Fistula in...
4 min · Published Feb 2020
Video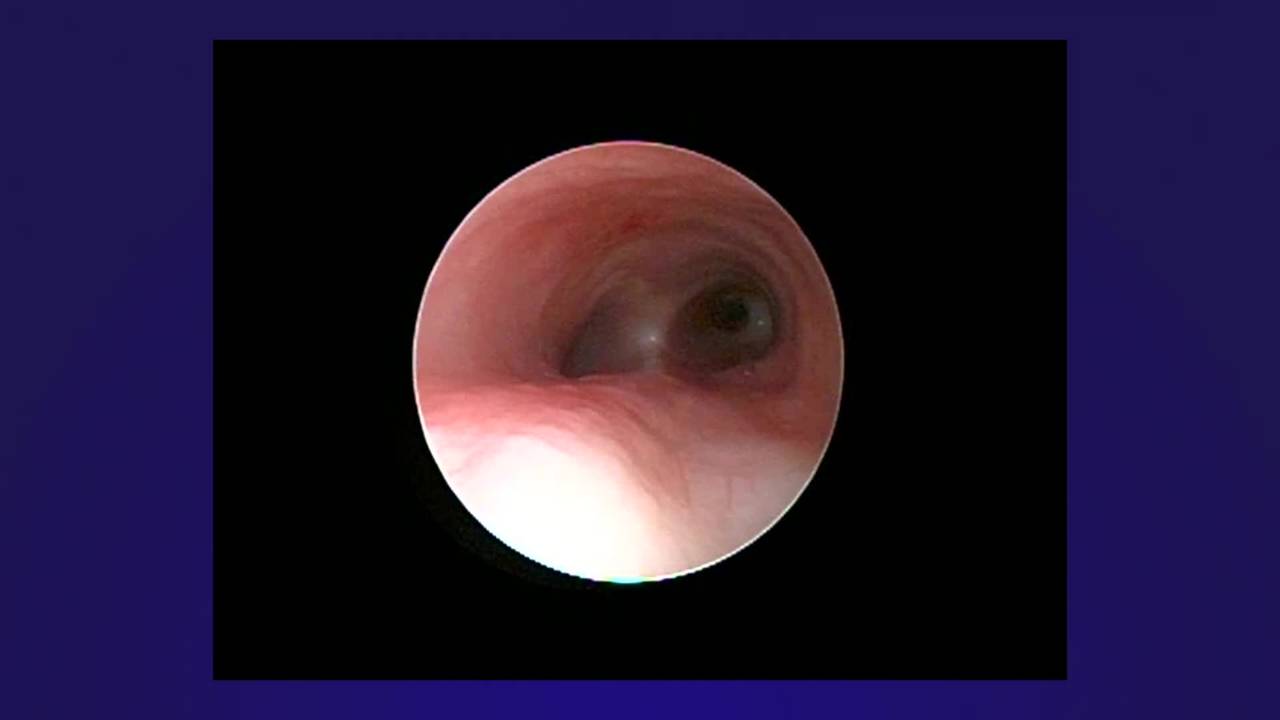
Rigid Bronchoscopy for Diagnosis and Treatment of Proximal Pouch Fistula in...
4 min · Published Feb 2020
Only a few other public items share this expert — go deeper there →
Video
What is Gastroschisis? An ERNICA animation for parents and families
2 min · Published Dec 2023
Video
What is Omphalocele? An ERNICA animation for parents and families
3 min · Published Dec 2023
Video
What is Congenital Diaphragmatic Hernia (CDH)? An ERNICA animation for parents and families
3 min · Published Dec 2023
Video
What is small bowel atresia?: An ERNICA animation for parents and families
4 min · Published Dec 2023
Video
Hirschsprung-associated enterocolitis in children: An ERNICA animation for parents and families
3 min · Published Dec 2023
Video
Surgical treatment for Hirschsprung’s Disease: An ERNICA animation for parents and families
3 min · Published Dec 2023
What the experts said
SEDA trial timeline: idea originated May 2017, ethical board approval January 2019, first patient included October 2019, 11th patient included 4 years after Rotterdam ethics approval
Reaching consensus on SEDA protocol took more than 1.5 years involving pediatric surgery, pediatrics, gastroenterology, pharmacy, endocrinology, and radiology departments
SEDA trial chose not to include patients younger than 3 months due to stricter regulations for young children, despite missing some rare patients
Ethical approval from Medical Review Ethics Committee can cost easily €2000 or more per center, regardless of whether study includes 500 or 10 patients
Clinical Trial Agreement (CTA) negotiations can take weeks to months, with one center taking almost 2 years before ethical approval could even start
SEDA trial budget of €750 per included patient covers research nurse, pharmacy, extra outpatient visits, additional esophagram, and travel costs, but not insurance, legal department, ethical approval, or translation costs
Different countries have different requirements: Sweden needs biobank approval for hair samples, Denmark has government-provided liability and subject insurance for all patients regardless of study participation
Centers with a research nurse or PhD candidate are more likely to complete ethical board and legal department processes compared to centers where full-time clinicians must take responsibility
MOUC-FAYA trial had more than 100 meetings in 18 months discussing idea, protocol versions, and statistical endpoints before first ethics approval
Primary endpoint of weight gain has many confounders including scale type, daytime of measurement, clothing, and edema, making it problematic for study results
MOUC-FAYA changed primary endpoint to time to full enteral feeds after enterostomic closure based on kilocalories per kilogram body weight in 24 hours, using standardized feeding protocol
Finding collaborating centers required more than 50 phone calls and several meetings at national and international conferences, resulting in 11 centers agreeing to participate
Grant application writing took 12 months, decision took 15 months; 80% of €800,000 German Research Foundation budget allocated to quality monitoring
MOUC-FAYA power analysis projected 120 patients (60 vs 60) with 201 patients expected for screening in 48 months (3 patients per center per year), but actual recruitment fell far behind with 62 patients recruited after extended timeline
Six MOUC-FAYA centers were excluded during study period due to poor recruitment; five new centers added including Austrian sites (Vienna, Graz) and planned Dutch sites (Amsterdam, Rotterdam)
MOUC-FAYA received second funding period adding €500,000 to original €800,000, totaling close to €1.5 million
Increasing protocol complexity negatively impacts patient recruitment and ability to maintain protocol adherence for data entry in electronic CRF
EU Clinical Trial Regulation became mandatory for all new medicinal product trial applications as of January 31, 2023 (two days before this webinar)
EU regulation differs from directive: regulation is Brussels-dictated with less room for national adaptation, while directives allow national legislation to add its own touch
EU Clinical Trial Regulation aims to address large decrease in clinical trials across European Union due to increasing costs, delayed startup, and lack of harmonization between member states
CTIS (Clinical Trial Information System) provides one single portal for submissions with one application dossier: Part I central (protocol, product information) and Part II national (patient information in local language, insurance, site assessment)
CTIS regulation mandates strict timelines for every process step; sponsors have only 10-12 days to respond to questions depending on stage, and failure to meet timelines can result in application lapse or tacit approval
Low-intervention clinical trials (LICT) qualify when: investigational medicinal product is authorized, used according to marketing authorization or as evidence-based treatment, and additional diagnostic/monitoring procedures pose no more than minimal risk or burden
For authorized medicinal products used on-label in non-blinded trials, additional labeling is not required and standard pharmacy prescription documentation can be used for traceability and accountability
Low-intervention clinical trials with minimal risk and burden can receive exemption from mandatory trial participant insurance (damage compensation), though liability insurance remains required
Clinical trial falls within EU regulation scope when studying medicinal product efficacy, safety, or pharmacokinetics/pharmacodynamics AND at least one of: therapeutic strategy assignment decided in advance outside normal practice, prescribing decision made simultaneously with trial inclusion decision, or additional diagnostic/monitoring procedures beyond normal practice applied
Randomization, blinding medication, administering products without marketing authorization, giving products without clinical indication, or having additional/more frequent diagnostic procedures all place trial outside normal clinical practice and within regulation scope
CTIS system is very complex with many roles and rights, strict timelines accounting for holidays across Europe, and transparency rules; recent German press publications raised concerns about postponing implementation due to system issues
Dutch CCMO offers pre-submission meetings to discuss best approach for submitting applications, available for trials to be performed in Netherlands
SEDA injection technique uses 1 mL triamcinolone (Kenalog) injected into 4 quadrants of esophagus at stenosis site using standard variceal injection needle (approximately 2mm depth) at 90 degrees or slightly oblique to create submucosal wheal
SEDA trial includes patients with type C esophageal atresia, age ≥3 months, at their 3rd dilatation to prevent refractory stricture (sometimes defined as ≥5 dilatations)
Penalties for violating clinical trial regulation differ between member states; in Netherlands, starting unauthorized trial or neglecting notification/result submission obligations can result in fines or jail time depending on seriousness
If sponsor application lapses due to missing 12-day timeline, there is a resubmission button in CTIS, but assessment fees increase; Netherlands has reduced fees for academic trials, Denmark has no fees, but fees vary by member state
For multinational trials, each participating member state charges its own fee—not just the lead member state—so fees multiply by number of countries
Sponsor is the entity (person or organization) responsible for initiation and conduct of clinical trial; in Dutch academic centers, typically the hospital board of directors is sponsor and appoints the principal investigator
For US or non-EU sponsors conducting trials in European Union, a legal representative of the sponsor must be appointed in EU with agreement on responsibilities; worldwide trials in rare diseases are common and feasible
Platform trial concept allows master protocol with initial submission explaining one arm, then subprotocols can be added via substantial modifications if within same hypothesis or primary objective
Substudies within main trial platform motivate centers by offering network for scientific collaboration using same cohort and existing biobank samples without adding additional trauma to patients
German Research Foundation limits non-German patients to less than 20% of total cohort in funded trials, creating obstacle for true European collaboration despite being most important German scientific funding organization
ERNICA research survey categorized European pediatric surgery research into four themes: origin, intervention, impact, and comorbidity